Monographs: Pharmaceutical substances: Amikacin (Amikacinum)
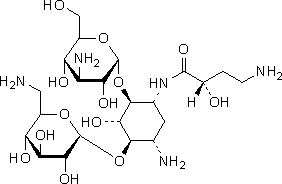
C22H43N5O13
Relative molecular mass. 585.6
Chemical name. 6-O-(3-amino-3-deoxy-α-D-glucopyranosyl)-4-O-(6-amino-6-deoxy-α-D-glucopyranosyl)-1-N-[(2S)-4-amino-2-hydroxybutanoyl]-2-deoxy-D-streptamine; 3-amino-3-deoxy-α-D-glucopyranosyl-(1→4)-[6-amino-6-deoxy-α-D-glucopyranosyl-(1→6)]-N3-[(2S)-4-amino-2-hydroxybutanoyl]-2-deoxy-L-streptamine; CAS Reg. No. 37517-28-5.
Description. White or almost white powder.
Solubility. Freely soluble in water, practically insoluble in acetone R or ethanol (~750 g/l) TS.
Category. Antibacterial.
Storage. Amikacin should be kept in a tightly closed container, or if sterile, in a hermetically closed container.
Labelling. The label states:
- where applicable, that the substances is free from bacterial endotoxins,
- where applicable, that the substance is sterile.
Requirements
Definition. Amikacin is a semi-synthetic product derived from a fermentation product, kanamycin A. Amikacin contains not less than 96.5% and not more than 102.0% (Assay, Method A) or not less than 98.5% and not more than 101.0% (Assay, Method B) of amikacin (C22H43N5O13), calculated with reference to the anhydrous substance.
Identity tests
• Either test A or tests B and C may be applied.
A. Carry out the test as described under 1.14.1 Chromatography, Thin-layer chromatography, using silica gel R5 as the coating substance and a mixture of 40 volumes of methanol R, 30 volumes of ammonia (~260 g/l) TS and 25 volumes of dichloromethane R as the mobile phase. Apply separately to the plate 2 µl of each of the following solutions in water R. For solution (A) use 5 mg of the test substance per mL. For solution (B) use 5 mg of amikacin RS per mL. For solution (C) use a mixture of 5 mg of amikacin RS and 5 mg of kanamycin monosulfate RS per mL. After removing the plate from the chromatographic chamber, heat it at 110 °C for 5 minutes, spray it with triketohydrindene/methanol reagent TS and heat further at 110 °C for 15 minutes.
The principal spot obtained with solution A corresponds in position, appearance, and intensity with that obtained with solution B. The test is not valid unless the chromatogram obtained with solution C shows two clearly separated spots.
B. Dissolve 10 mg in 1 mL of water R, add 1 mL of sodium hydroxide (~80 g/l) TS and mix, then add 2 mL of cobalt(II) nitrate (10 g/l) TS; a violet colour with precipitate is produced.
C. Dissolve 0.05 g in 3 mL of water R and add 4 mL of anthrone TS; a bluish violet colour is produced.
Specific optical rotation (1.4). Use a 20 mg/mL solution and calculate with reference to the anhydrous substance:  = +97° to +105°.
Sulfated ash (2.3). After ignition moisten the residue with 2 mL of nitric acid (~1000 g/l) TS and about 0.2 mL of sulfuric acid (~1760 g/l) TS; not more than 5 mg/g.
Water. Determine as described under 2.8 Determination of water by the Karl Fischer method, Method A, using about 0.2 g of the substance; the water content is not more than 85 mg/g.
pH value (1.13). pH of a 10 mg/mL solution in carbon-dioxide-free water R, 9.5-11.5.
Bacterial endotoxins. If intended for use in the manufacture of a parenteral dosage form, carry out the test as described under 3.4 Test for bacterial endotoxins; contains not more than 0.33 IU of endotoxin per mg of amikacin.
Sterility. If intended for use in the manufacture of either a parenteral or other sterile dosage form without a further appropriate sterilization procedure, complies with 3.2 Test for sterility.
Kanamycin A. Carry out the test as described under 1.14.1 Chromatography, Thin-layer chromatography, using silica gel R5 as the coating substance and a mixture of 40 volumes of methanol R, 30 volumes of ammonia (~260 g/l) TS and 25 volumes of dichloromethane R as the mobile phase. Apply separately to the plate 5 µl of each of the following solutions in water R. For solution (A) use 40 mg of the test substance per mL. For solution (B) use 1.6 mg of kanamycin monosulfate RS per mL. For solution (C) use a mixture of 2 mg of amikacin RS and 2 mg of kanamycin monosulfate RS per mL. After removing the plate from the chromatographic chamber, heat it at 110 °C for 5 minutes, spray it with triketohydrindene/methanol reagent TS and heat further at 110 °C for 15 minutes.
In the chromatogram obtained with solution A, any spot corresponding to kanamycin A, is not more intense than that obtained with solution B (4.0%). The test is not valid unless the chromatogram obtained with solution C shows two clearly separated spots.
Related substances. Carry out the test as described under 1.14.1 Chromatography, High-performance liquid chromatography, using a stainless steel column (25 cm x 4.6 mm) packed with base-deactivated particles of silica gel the surface of which has been modified with chemically bonded octadecylsilyl groups (5 µm).
The mobile phases for gradient elution consist of a mixture of Mobile phase A and Mobile phase B, using the following conditions:
Mobile phase A: dissolve 1.8 g of sodium octanesulfonate R and 20.0 g of anhydrous sodium sulfate R with 50.0 mL of 0.2 M phosphate buffer pH 3.0 and 50.0 mL of acetonitrile R. Dilute to 1000 mL with water R.
Mobile phase B: dissolve 1.8 g of sodium octanesulfonate R and 20.0 g of anhydrous sodium sulfate R with 50.0 mL of 0.2 M phosphate buffer pH 3.0 and 100.0 mL of acetonitrile R. Dilute to 1000 mL with water R.
Prepare the 0.2 M phosphate buffer pH 3.0 by mixing 0.2 M potassium dihydrogen phosphate R with 0.2 M phosphoric acid R until pH 3.0 is reached.
|
Time |
Mobile phase A |
Mobile phase B |
Comments |
|
0-30 |
100 |
0 |
Isocratic |
|
30-75 |
100 to 0 |
0 to 100 |
Linear gradient |
|
75-77 |
0 to 100 |
100 to 0 |
Return to initial composition |
|
77-87 |
100 |
0 |
Re-equilibration |
Prepare the following solutions in mobile phase A. For solution (1) dissolve 50.0 mg of the test substance in 10.0 mL. For solution (2) dilute 1.0 mL of solution (1) to 100.0 mL.
Operate with a flow rate of 1.0 mL per minute. As a detector use an ultraviolet spectrophotometer set at a wavelength of 200 nm. Maintain the temperature of the column at 40°C.
Inject separately 20 µl each of solutions (1) and (2). The test is not valid unless in the chromatogram obtained with solution (2) the retention time of the principal peak is between 25 and 30 minutes and the signal-to-noise ratio is at least 20. When the retention time of the principal peak is not reached, adjust the content of acetonitrile R in mobile phase A. In the chromatogram obtained with solution (1), the area of any peak, other than the principal peak, is not greater than twice the area of the principal peak obtained with solution (2) (2.0%). The sum of the areas of all peaks, other than the principal peak is not greater than four times the area of the principal peak obtained with solution (2) (4.0%). Disregard any peak with an area less than 0.5 times the area of the principal peak obtained with solution (2) (0.5%).
Assay
• Either method A or B may be applied.
A. Carry out the assay as described under 1.14.1 Chromatography, High-performance liquid chromatography, using a stainless steel column (25 cm x 4.6 mm) packed with base-deactivated particles of silica gel the surface of which has been modified with chemically bonded octadecylsilyl groups (5 µm).
As the mobile phase use a solution prepared as follows: dissolve 1.8 g of sodium octanesulfonate R and 20.0 g of anhydrous sodium sulfate R with 50.0 mL of 0.2 M phosphate buffer pH 3.0 and 50.0 mL of acetonitrile R. Dilute to 1000 mL with water R.
Prepare the 0.2 M phosphate buffer pH 3.0 by mixing 0.2 M potassium dihydrogen phosphate R with 0.2 M phosphoric acid R until pH 3.0 is reached.
Prepare the following solutions in the mobile phase. For solution (1) dissolve 50.0 mg of the test substance in 25.0 mL. For solution (2) dissolve 50.0 mg of amikacin RS in 25.0 mL.
Operate with a flow rate of 1.0 mL per minute. As a detector use an ultraviolet spectrophotometer set at a wavelength of 200 nm. Maintain the temperature of the column at 40°C.
Inject separately 20 µl each of solutions (1) and (2). The assay is not valid unless the symmetry factor of the principal peak is less than 1.5.
Measure the areas of the peak responses obtained in the chromatograms from solutions (1) and (2), and calculate the content of amikacin (C22H43N5O13).
B. Dissolve about 0.1 g of the test substance, accurately weighed, in 30 mL of glacial acetic acid R1, and titrate with perchloric acid (0.1 mol/l) VS as described under 2.6 Non-aqueous titration, Method A. Each mL of perchloric acid (0.1 mol/l) VS is equivalent to 14.64 mg of C22H43N5O13.
Impurities
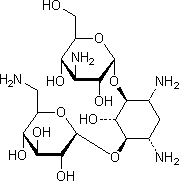
A. 4-O-(3-amino-3-deoxy-α-D-glucopyranosyl)-6-O-(6-amino-6-deoxy-α-D-glucopyranosyl)-2-deoxy-L-streptamine (kanamycin A).
 |  |